make preferable
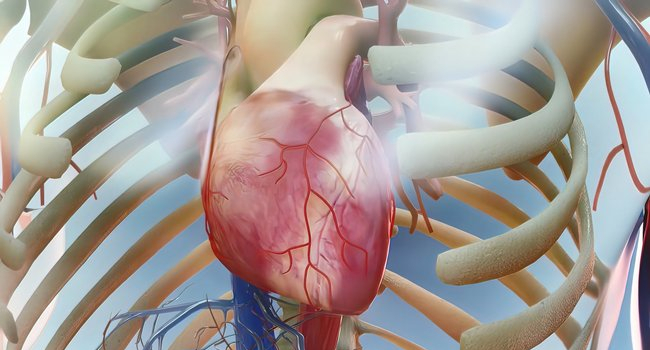
Researchers from the Carlos III National Center for Cardiovascular Research (CNIC), together with an international team of scientists, have identified a new molecular mechanism involved in the development of hypertrophic cardiomyopathy (HCM), the most common inherited heart disease. The findings were published in the journal Nature Cardiovascular Research.
Hypertrophic cardiomyopathy is one of the leading causes of sudden cardiac death in young people and athletes. According to epidemiological studies, the condition affects approximately one in every 250–500 individuals in the general population. In Spain alone, around 95,000 people are living with the disease.
The disorder is characterized by abnormal thickening of the heart muscle and excessive contraction of cardiomyocytes—the cells responsible for generating the force of each heartbeat. As a result, blood flow out of the heart may become obstructed, while overly forceful contractions can trigger life-threatening heart rhythm disturbances.
In most cases, HCM is caused by mutations in genes encoding sarcomeric proteins—the molecular machinery responsible for heart muscle contraction. One of the most commonly affected genes is MYBPC3, which encodes cardiac myosin-binding protein C (cMyBP-C).
The new study sheds light on how one specific group of MYBPC3 mutations causes disease. According to the study's first author, Laura San-Martín, unlike the most common mutations, this variant does not reduce the amount of cMyBP-C in heart cells. Instead, it disrupts the protein's ability to interact with other cardiac proteins.
To investigate this mechanism, the researchers developed a mouse model carrying the R502W mutation in the cMyBP-C protein. Their analysis revealed that the mutation weakens the protein's ability to bind to myosin—the molecular "motor" responsible for heart muscle contraction. This impaired interaction represents a previously unknown disease mechanism affecting a subset of patients with hypertrophic cardiomyopathy.
The team also evaluated the effects of mavacamten, the first drug specifically developed to treat hypertrophic cardiomyopathy. Mavacamten targets myosin, regulating its activity and reducing excessive contraction of the heart muscle.
The experiments demonstrated that treatment prevented pathological remodeling of the heart in both mice carrying the R502W mutation and in models with complete loss of cMyBP-C. In addition, mice with the R502W mutation showed improved exercise tolerance. The therapeutic effect was also confirmed in laboratory-grown human heart tissue derived from induced pluripotent stem cells.
The researchers conclude that the effectiveness of mavacamten appears to be independent of the specific mutation responsible for the disease. This finding broadens the range of patients who could potentially benefit from the therapy.